


When a molecule absorbs light from its surroundings, this can induce molecular motion, change the electronic structure, eject electrons, or re-emit the light (and often a combination of the above). Microwaves excite rotational motion and are used by microwave ovens to heat your food. Infrared induces vibrational motion, and is responsible for the sun’s warmth. Visible and UV light induce electronic excitations which are responsible for the color of dyes, the activity of solar cells, and the degradation of polymers. X-rays can rip core electrons out, generating radicals that can cause DNA damage.
Rowan has long supported the prediction of molecular vibrations and the resulting IR spectra, but the computation of electronic excitations is significantly more involved and requires excited-state methods. We are excited to introduce TDDFT on Rowan, providing access to UV-Vis absorbance and fluorescence prediction as well as excited-state geometry optimization. For more on the specifics of how TDDFT works, you can read our technical blog on the method. For a worked example of using TDDFT to predict the color of azo dyes, you can read a blog post by our summer intern Isaiah.
Absorbance
When light of the correct energy hits a molecule, it can excite the molecule to a new electronic state wherein the electron configuration differs. Once excited, the light may be immediately re-emitted at the same energy (Rayleigh scattering), re-emitted with a slight shift (Raman scattering), or the molecule can remain in the excited state for an extended period of time and change geometry (potentially emitting from the new geometry). The wavelengths at which a molecule absorbs can be predicted with TDDFT.
Rowan runs TDDFT with GPU4PySCF on H200 GPUs, providing significant speedups over CPU-based implementations. To run TDDFT starting from an optimized ground-state geometry:
Open the basic calculation workflow, and select the TDDFT mode.
Select single-point energy (the “Energy” task).
Pick a functional, basis set, and solvent; ωB97X-D3/def2-TZVP is a great place to start.
Choose how many roots to compute; we recommend 10 for small molecules (<30 atoms) and 20–50 for larger molecules.
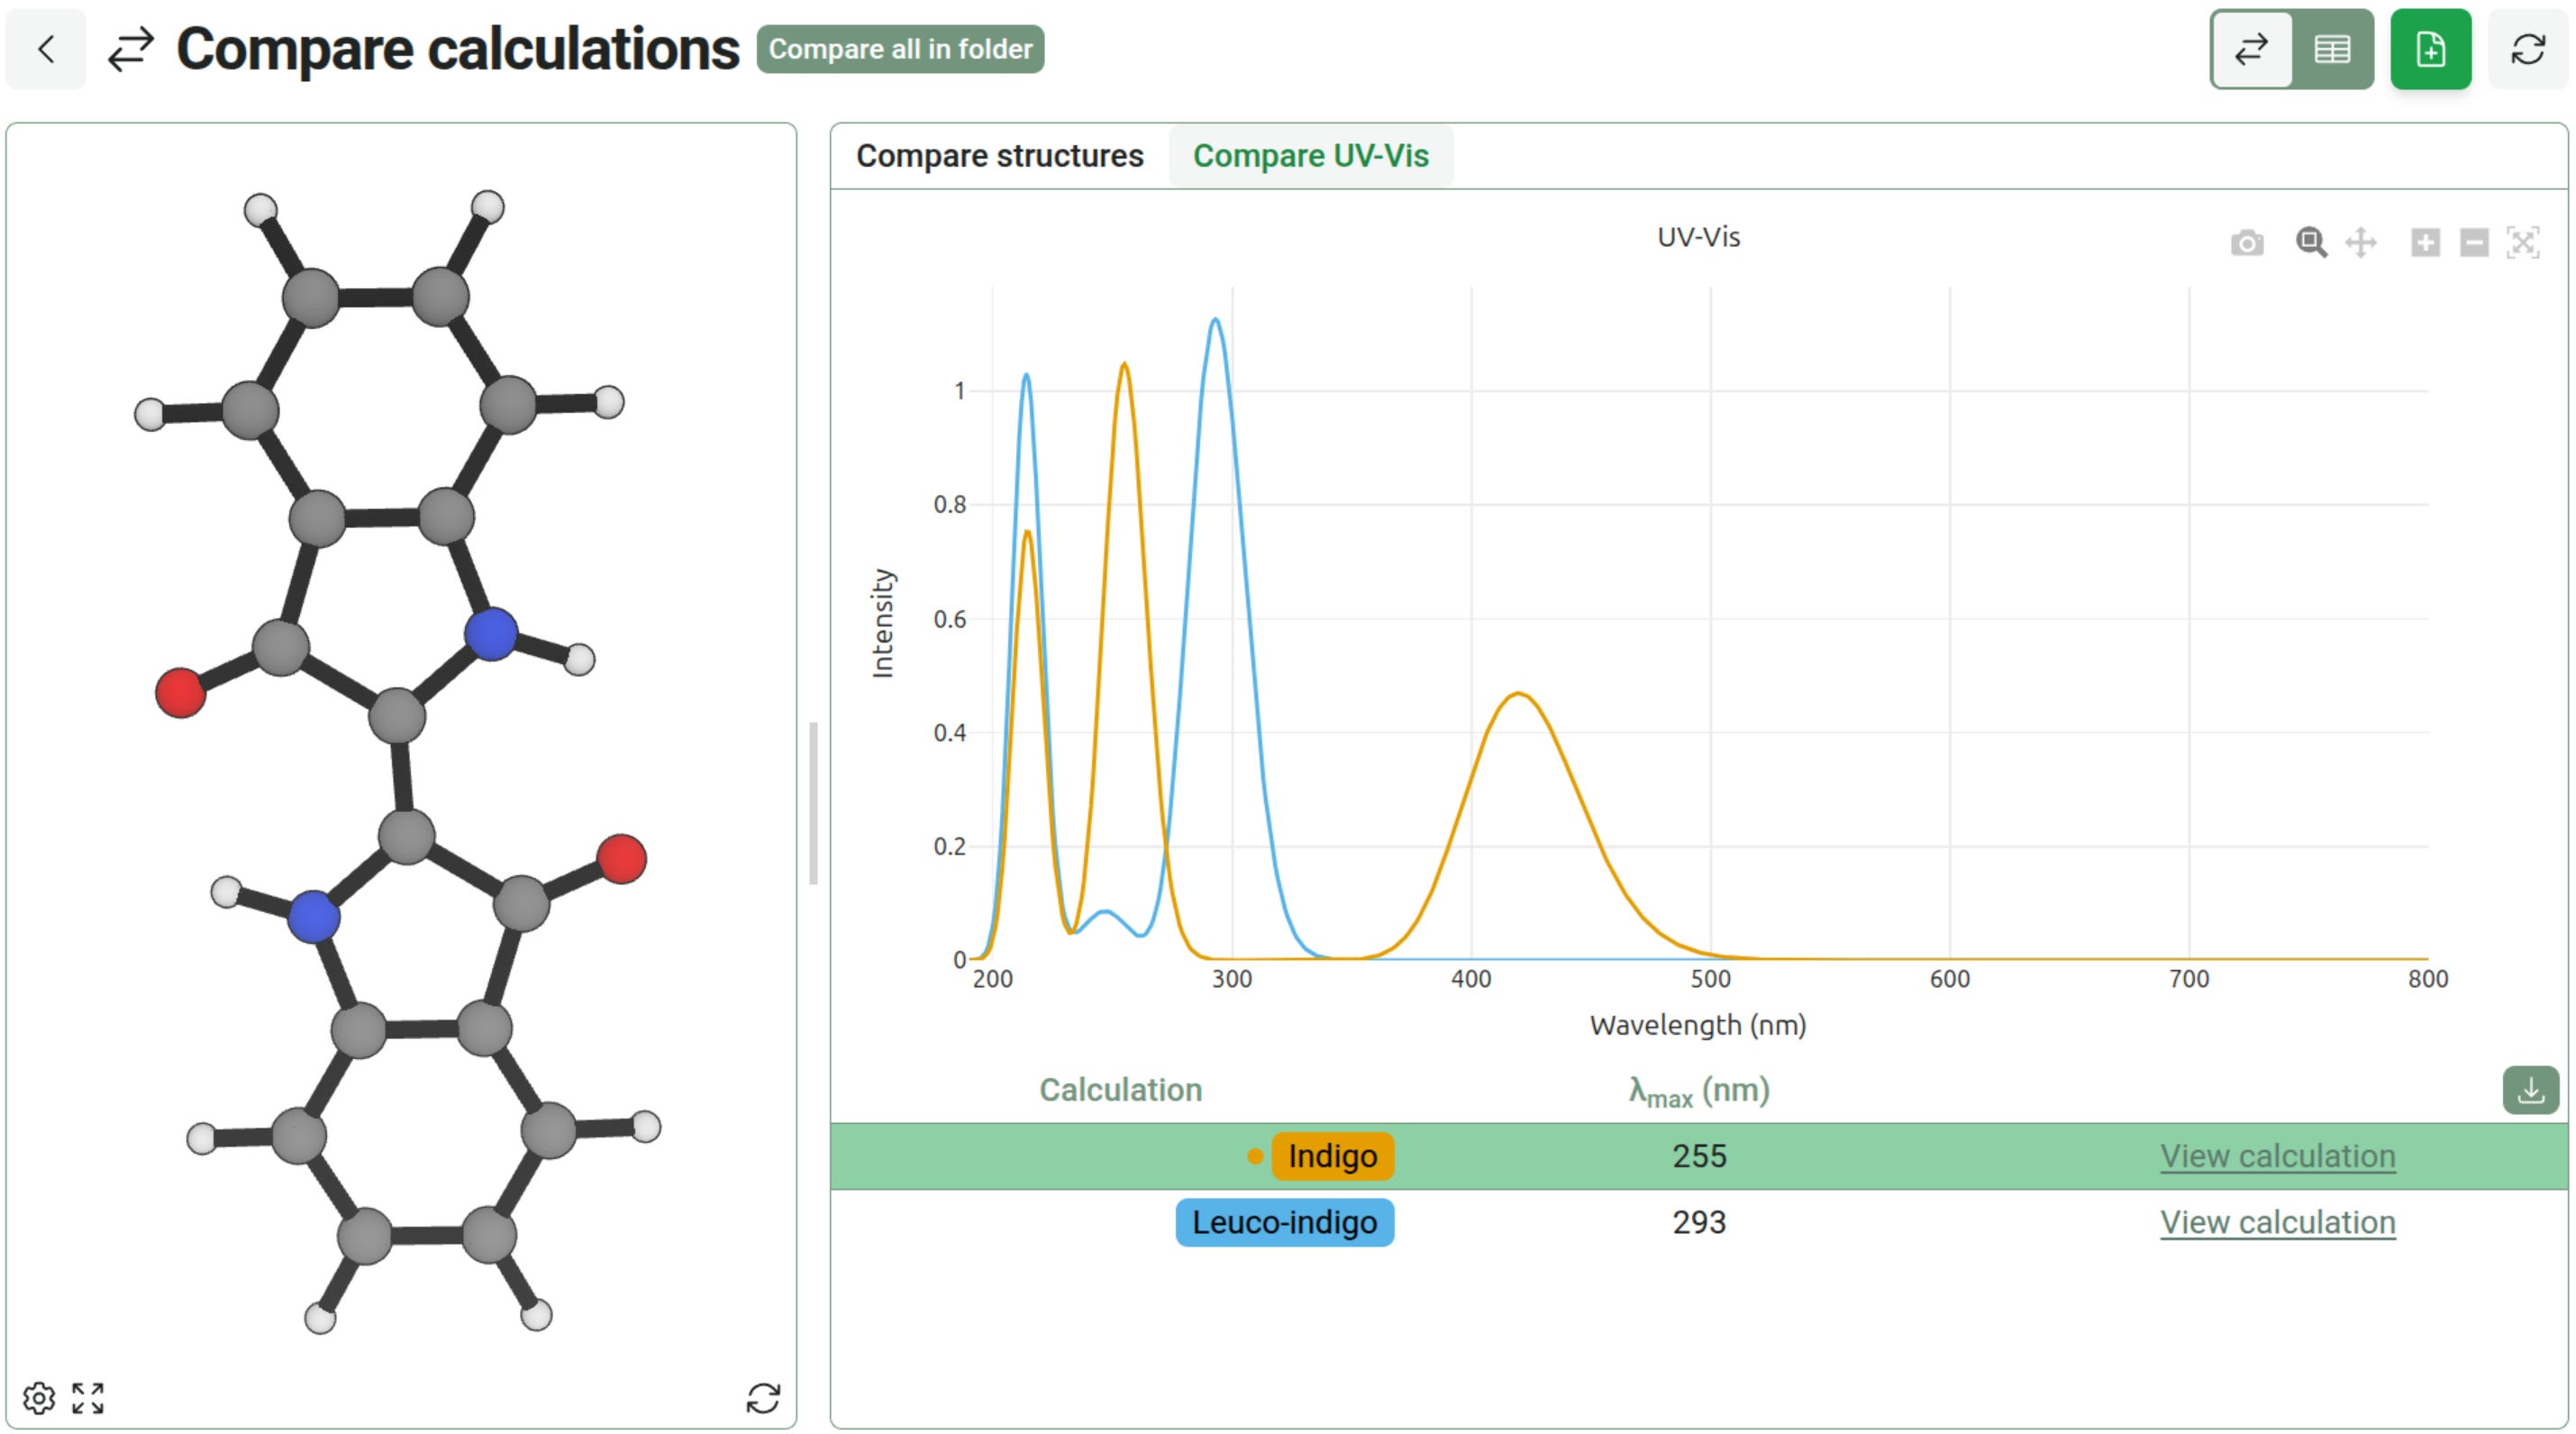
Molecules that absorb in the visible range can be useful as dyes, and their color is dependent on the visible light that the molecule does not absorb. Single-point TDDFT calculations can provide a useful approximation of shifts in the absorbance peak (λmax) when modifying dyes, but care should be taken when reporting absolute λmax values (see an in-depth way of handling this for a large number of dyes in Isaiah’s blog post). Errors in λmax are often due to a combination of:
XC functional error,
lack of vibronic effects,
self-interaction error,
the adiabatic approximation, and
solvation effects.
If you are interested in running more accurate UV-Vis absorbance calculations at scale, we’d be happy to chat about the ways to improve your calculations; please reach out.
Fluoresence
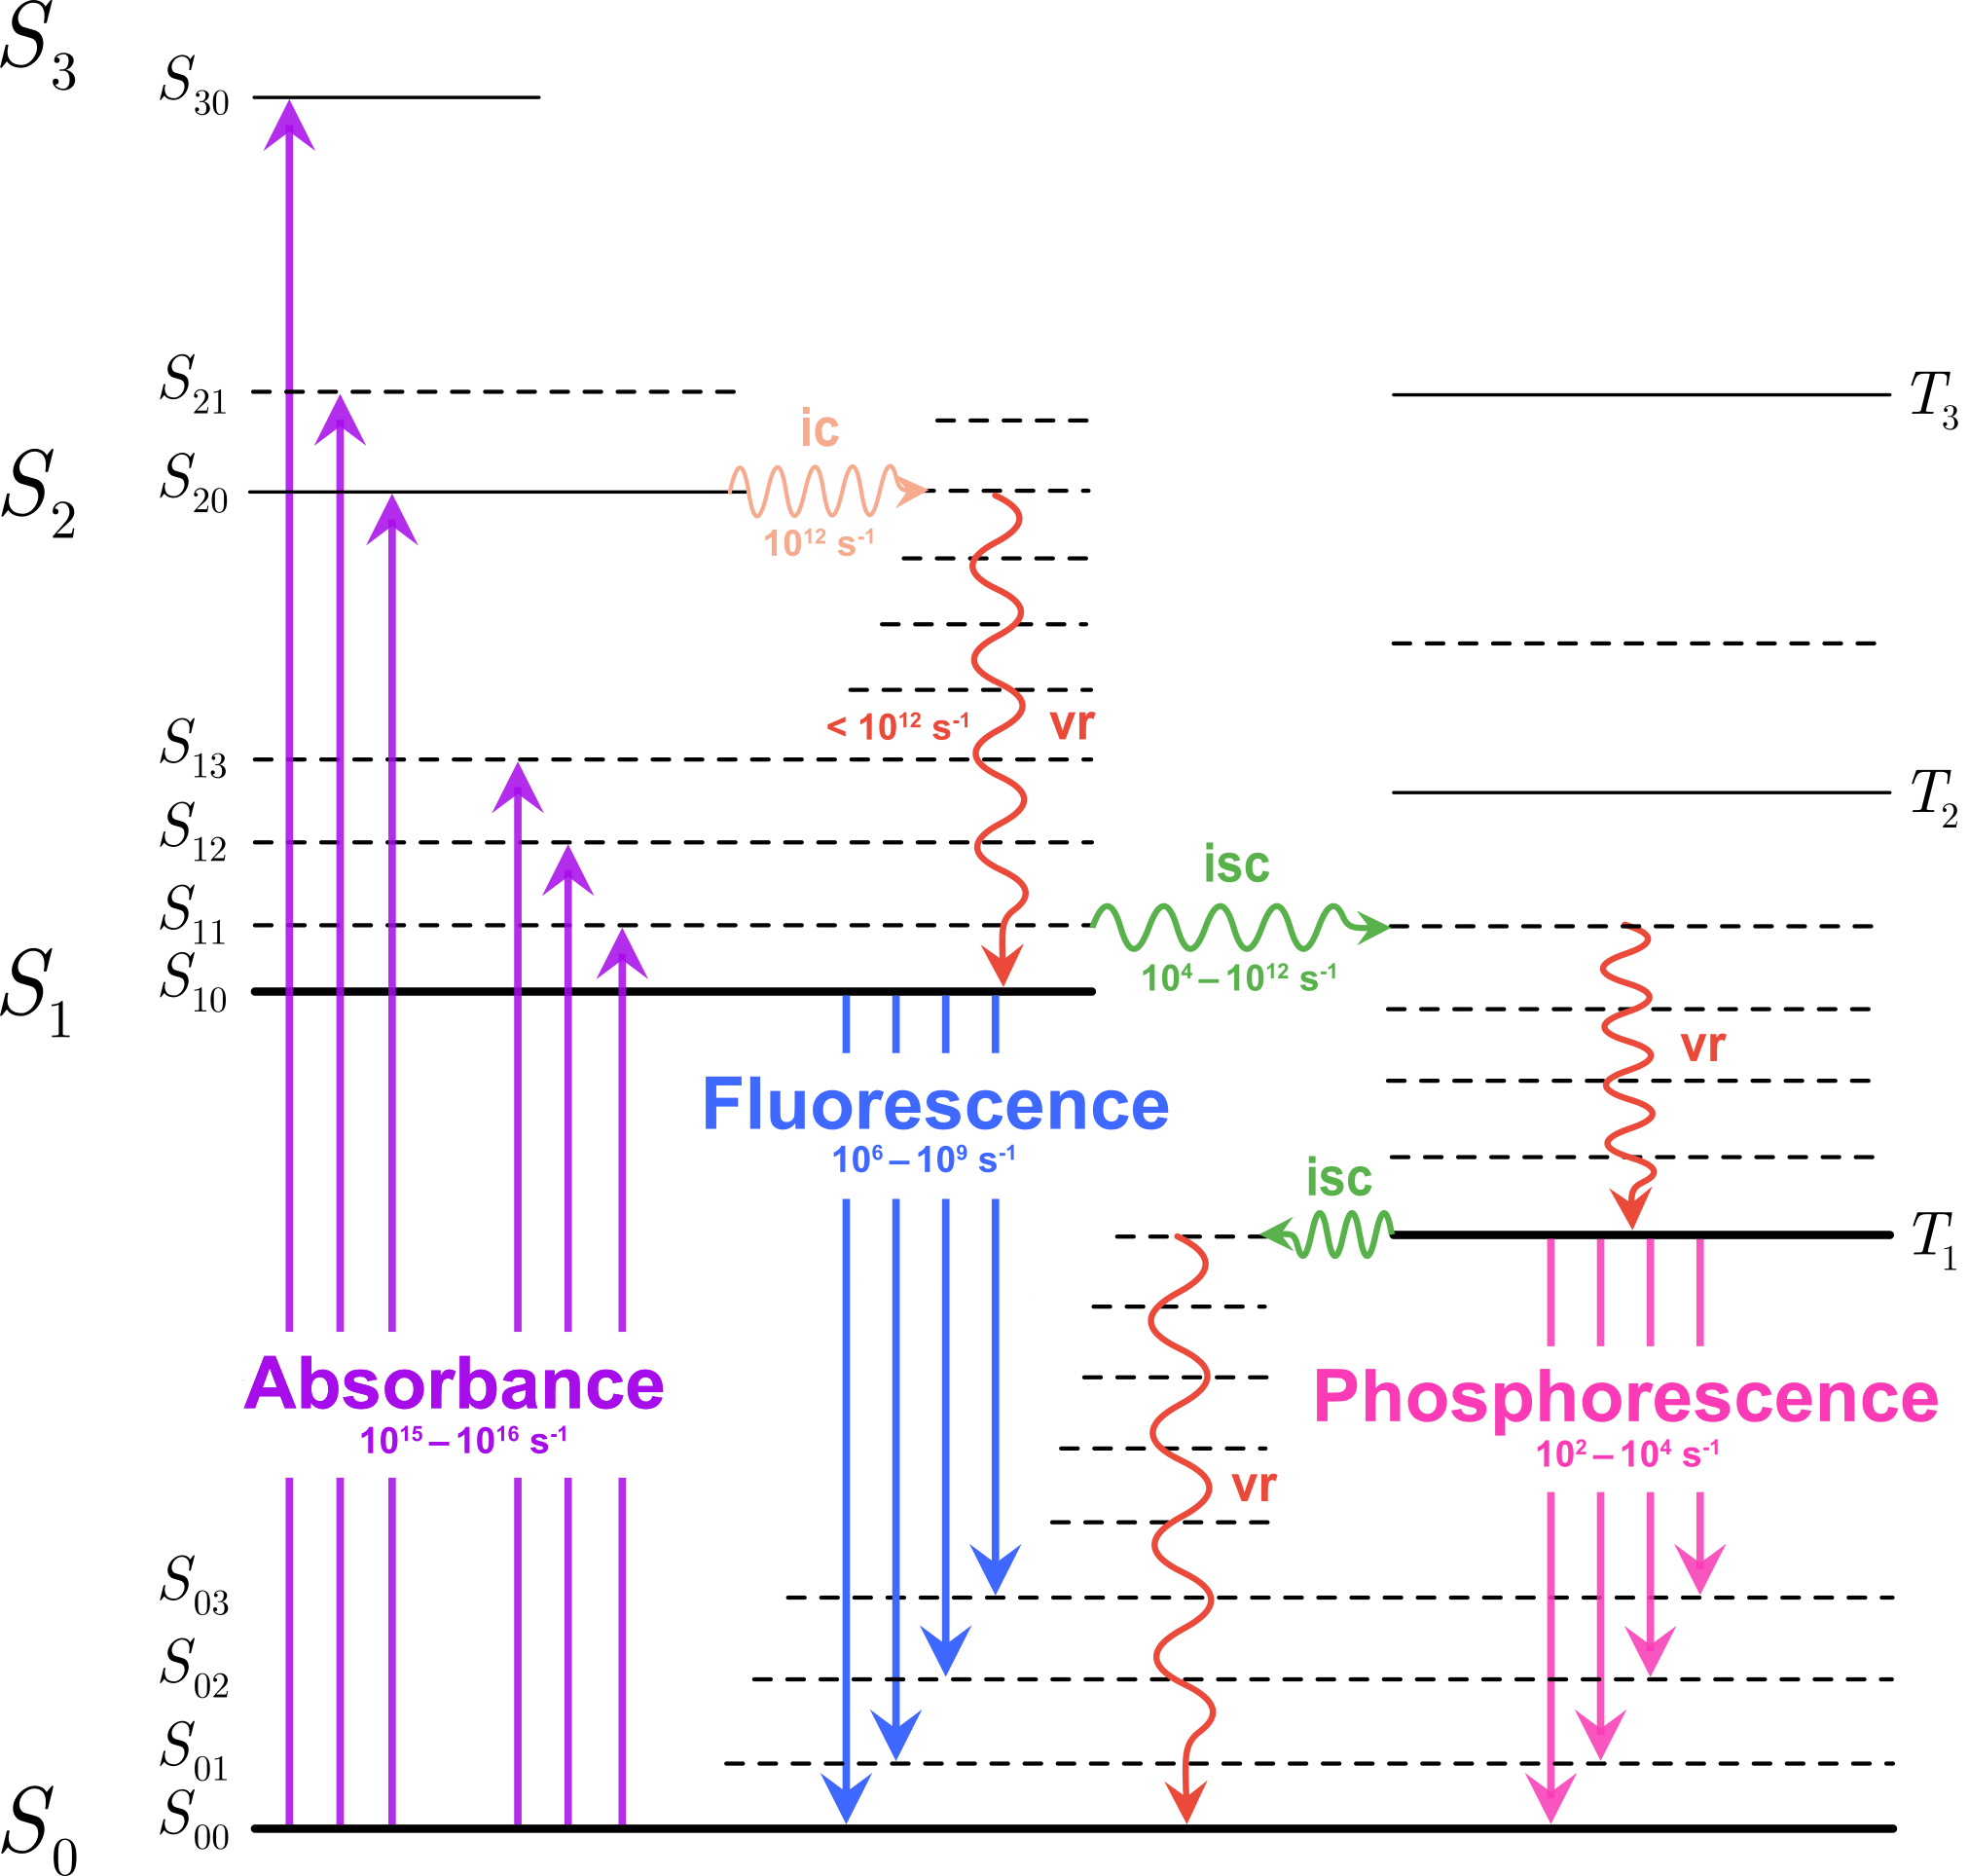
When molecules are first excited, they are at a non-optimal geometry on the excited-state surface. If the light is not immediately re-emitted, vibrational relaxation quickly occurs (<10–12 seconds), producing a new geometry. When the molecule fluoresces from the new geometry (10–6–10–9 seconds) and drops down to the ground electronic state, it emits less energy than it absorbed since it lands on a different section of the ground-state potential-energy surface (observed as a Stokes shift in fluorescence spectroscopy).
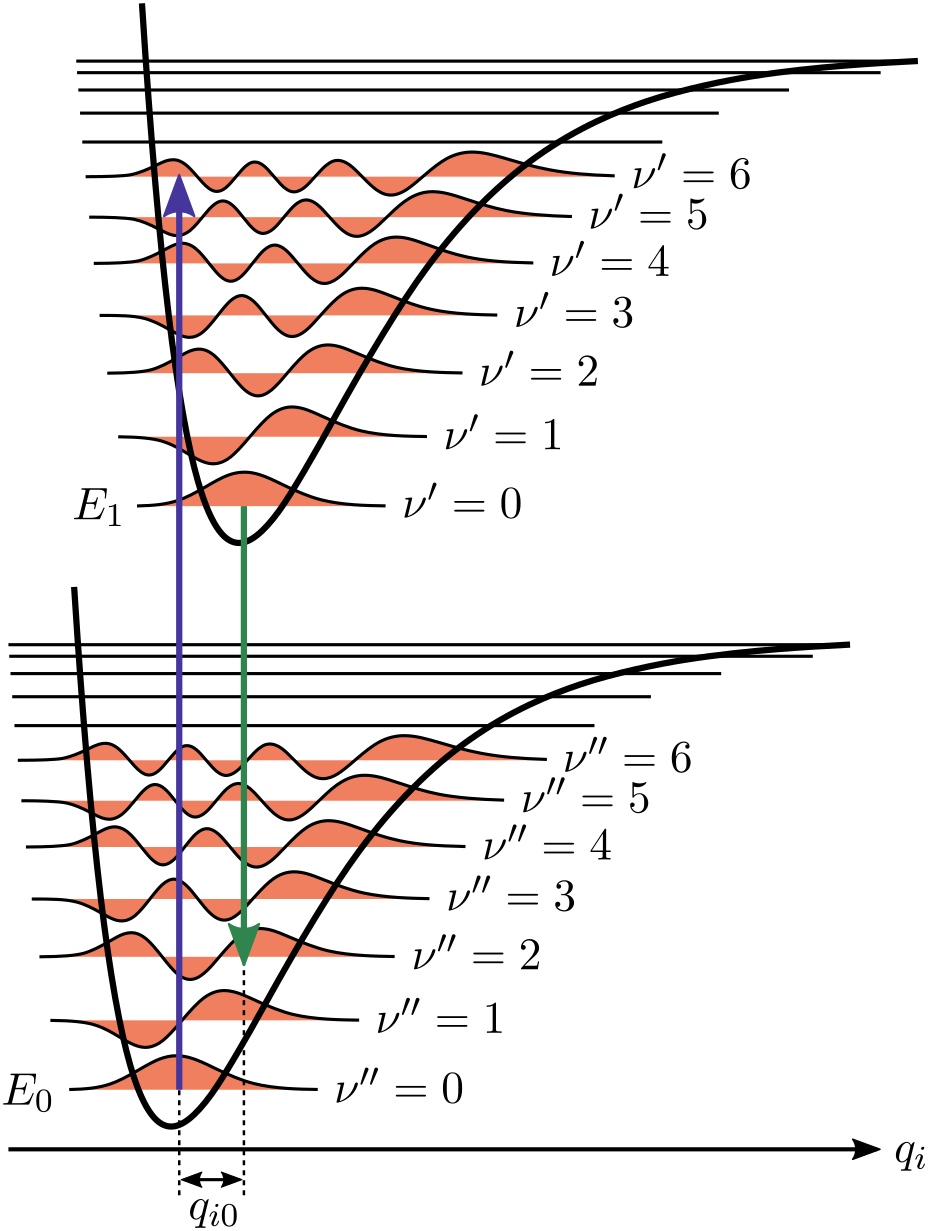
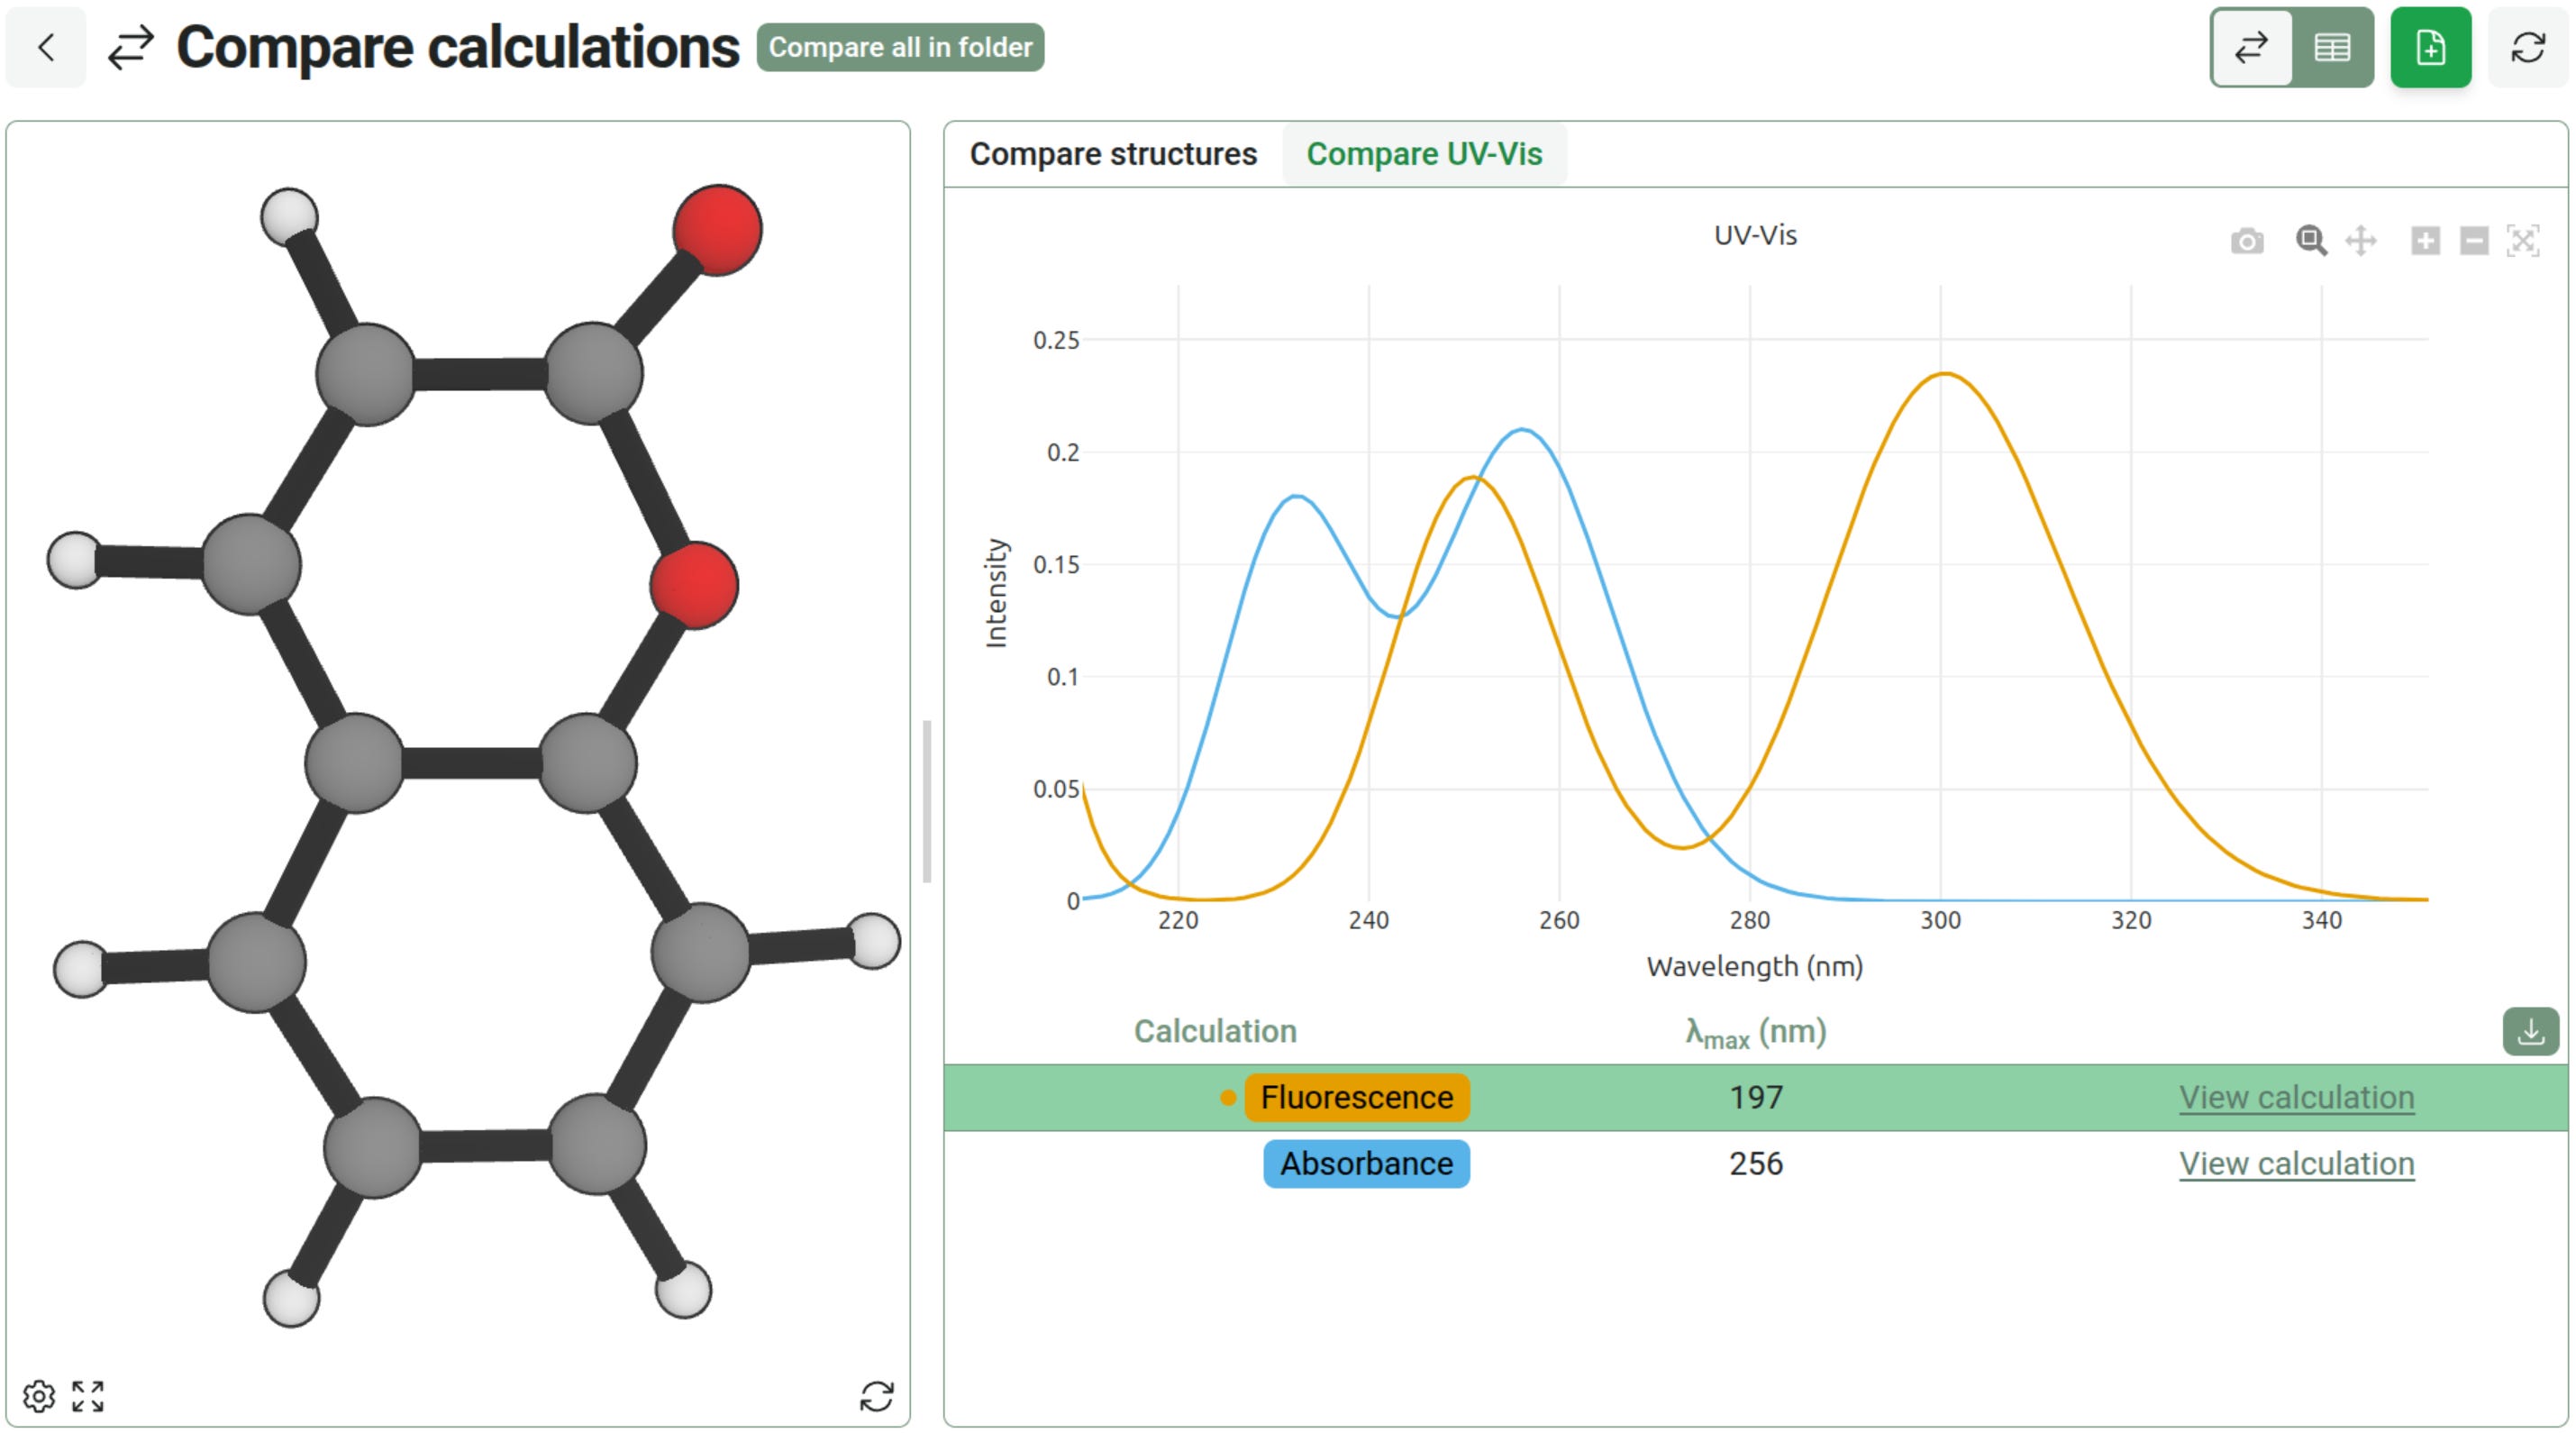
Fluorescence is significantly harder to model than absorbance due to the variable excited state lifetimes, which are affected by the rates of:
Internal conversions (IC) - conversion between electronic states of the same multiplicity
Intersystem crossing (ISC) - conversion between electronic states of the different multiplicity
Vibrational relaxation (VR) - relaxation of vibrationally excited (“hot”) molecules down to their lowest vibrational state
Phosphorescence - emission from an excited state of a different multiplicity than the ground state
Internal conversion and vibrational relaxation are typically fast compared to fluorescence, and intersystem crossing is strongly dependent on the potential for spin-orbit coupling, which is more pronounced in heavier elements. Vibrational relaxation down to the ground state can occur through intersystem crossing, avoiding the fluorescence path entirely.
Light-induced rearrangements
When molecules relax after being excited to a new electronic state, significant rearrangement can be induced. Tetrafluoroethylene’s geometry changes from flat (D2h) to twisted (D2d) in its first excited state; similar rotations cause the cis–trans isomerizations that allow retinal in our eyes to detect light. Photoacids like triphenylsulfonium triflate break internal bonds and rearrange to produce a strong acid upon irradiation. Photochromic species shift the wavelengths of light they can absorb, allowing for things like photochromic glasses to act as sunglasses outside. Polymers with significant unsaturation will often produce radicals under UV irradiation, leading to a radical cascade and photodegradation; this leads to the yellowing of styrene–butadiene paints and the need for photostabilizers.
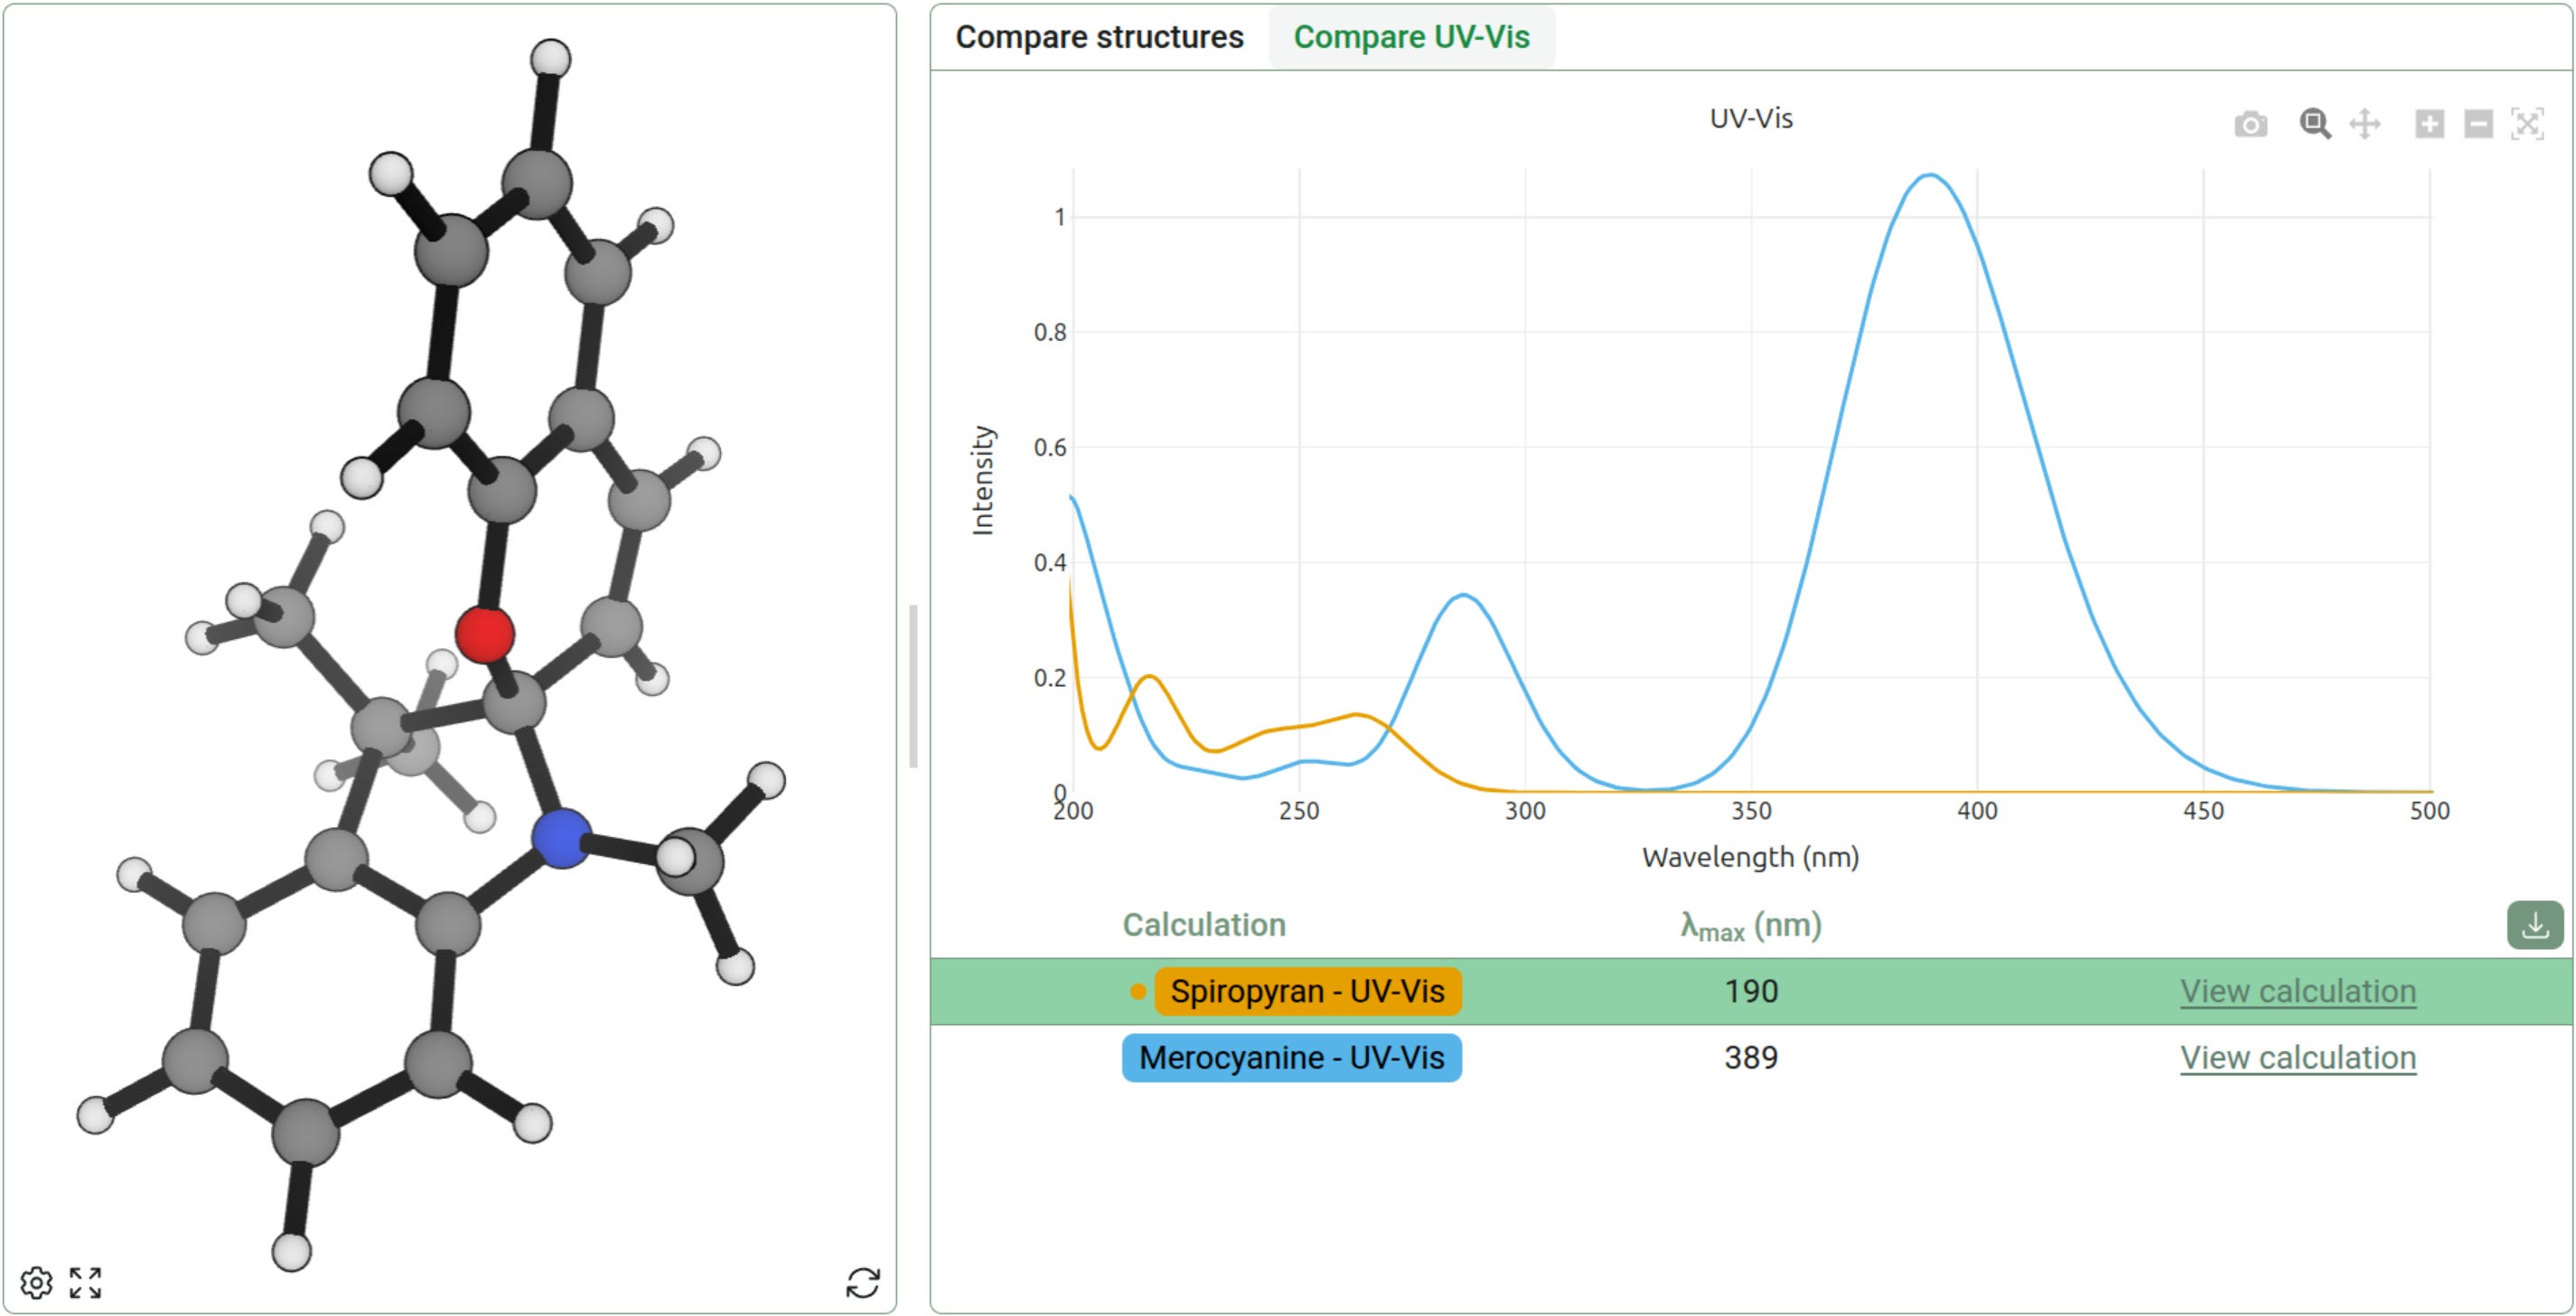
In spiropyrans, excitation by UV light leads to C–O bond cleavage and a flattening of the species into a merocyanine. This can be modeled via optimization of the first excited state with TDDFT. Once the excited-state optimized species relaxes back to the ground state, merocyanine is formed. This species has significantly more absorbance in the visible range due to its extended π system. Modified spiropyrans can shift the absorption wavelength to different colors, and can be used as photochromic indicators.
Conclusion
TDDFT can be used to model numerous systems beyond what has already been shown, including:
Photodegradation of polymers
OLED emission spectra, degradation, and quenching
Solar cell sensitivity
Biomolecular UV response
You can model all of these today on the Rowan platform. Since TDDFT is significantly slower than ground-state DFT, we recommend subscribing if you plan to run more than a couple UV-Vis curves or if you plan to run excited-state optimizations. TDDFT is available in both our API and via the GUI, and the latter is great for large-scale screening.
If you are interested in running calculations at scale, running more accurate excited-states methods like EOM-CCSD, or modeling advanced properties like circular dichroism, please get in touch; we’d be happy to chat.


